How-to guides
Pick a task. Follow the numbered steps. Try it opens the live view. Click a step number to mark it done.
Which tool do I need?
Filter dsRNAs
All 5.1 million predictions as a searchable table, with the same confidence filters as IGV.
- Search by gene symbols, Ensembl IDs or a locus
- Filter by model scores, pairing, length, energy, conservation, editing
- Summary counts, distributions, downloads (CSV, Parquet, BEDPE)
IGV Viewer
Predictions drawn as arcs on the genome, with gene models, repeats, editing sites, conservation and RBP binding.
- Jump to a gene or coordinates
- Click an arc for scores and a link to its structure
- Share links that restore your locus, tracks and filters
dsRNA Browser
One dsRNA at a time: predicted structure, chemical probing reactivity, editing sites, an RNAfold alternative, and a downloadable package.
- A selected set of ~108,000 dsRNAs that overlap exons (where probing data exist); 13,789 of them have probing coverage that passes the power filter
- ROC-AUC per arm and per experiment, with confidence intervals
- Printable summary and ZIP of FASTA / DBN / CT / BED / SVG
Downloads
The full dataset, browser track files, a column dictionary, and recipes for pandas / polars / DuckDB and bedtools.
- Full dsRNAome as Parquet and BEDPE
- IGV track files for your own IGV desktop session
- Version and changelog
Typical path:
Walkthroughs
Examples use ZNF426 (chr19; a long, nearly perfect dsRNA in its 3′ UTR), MAVS (chr20) and SON (chr21, the IGV default).
1. Look at the dsRNAs in your gene
IGV ViewerEvery predicted dsRNA in a gene, next to exons, repeats, editing sites and conservation.
-
Type
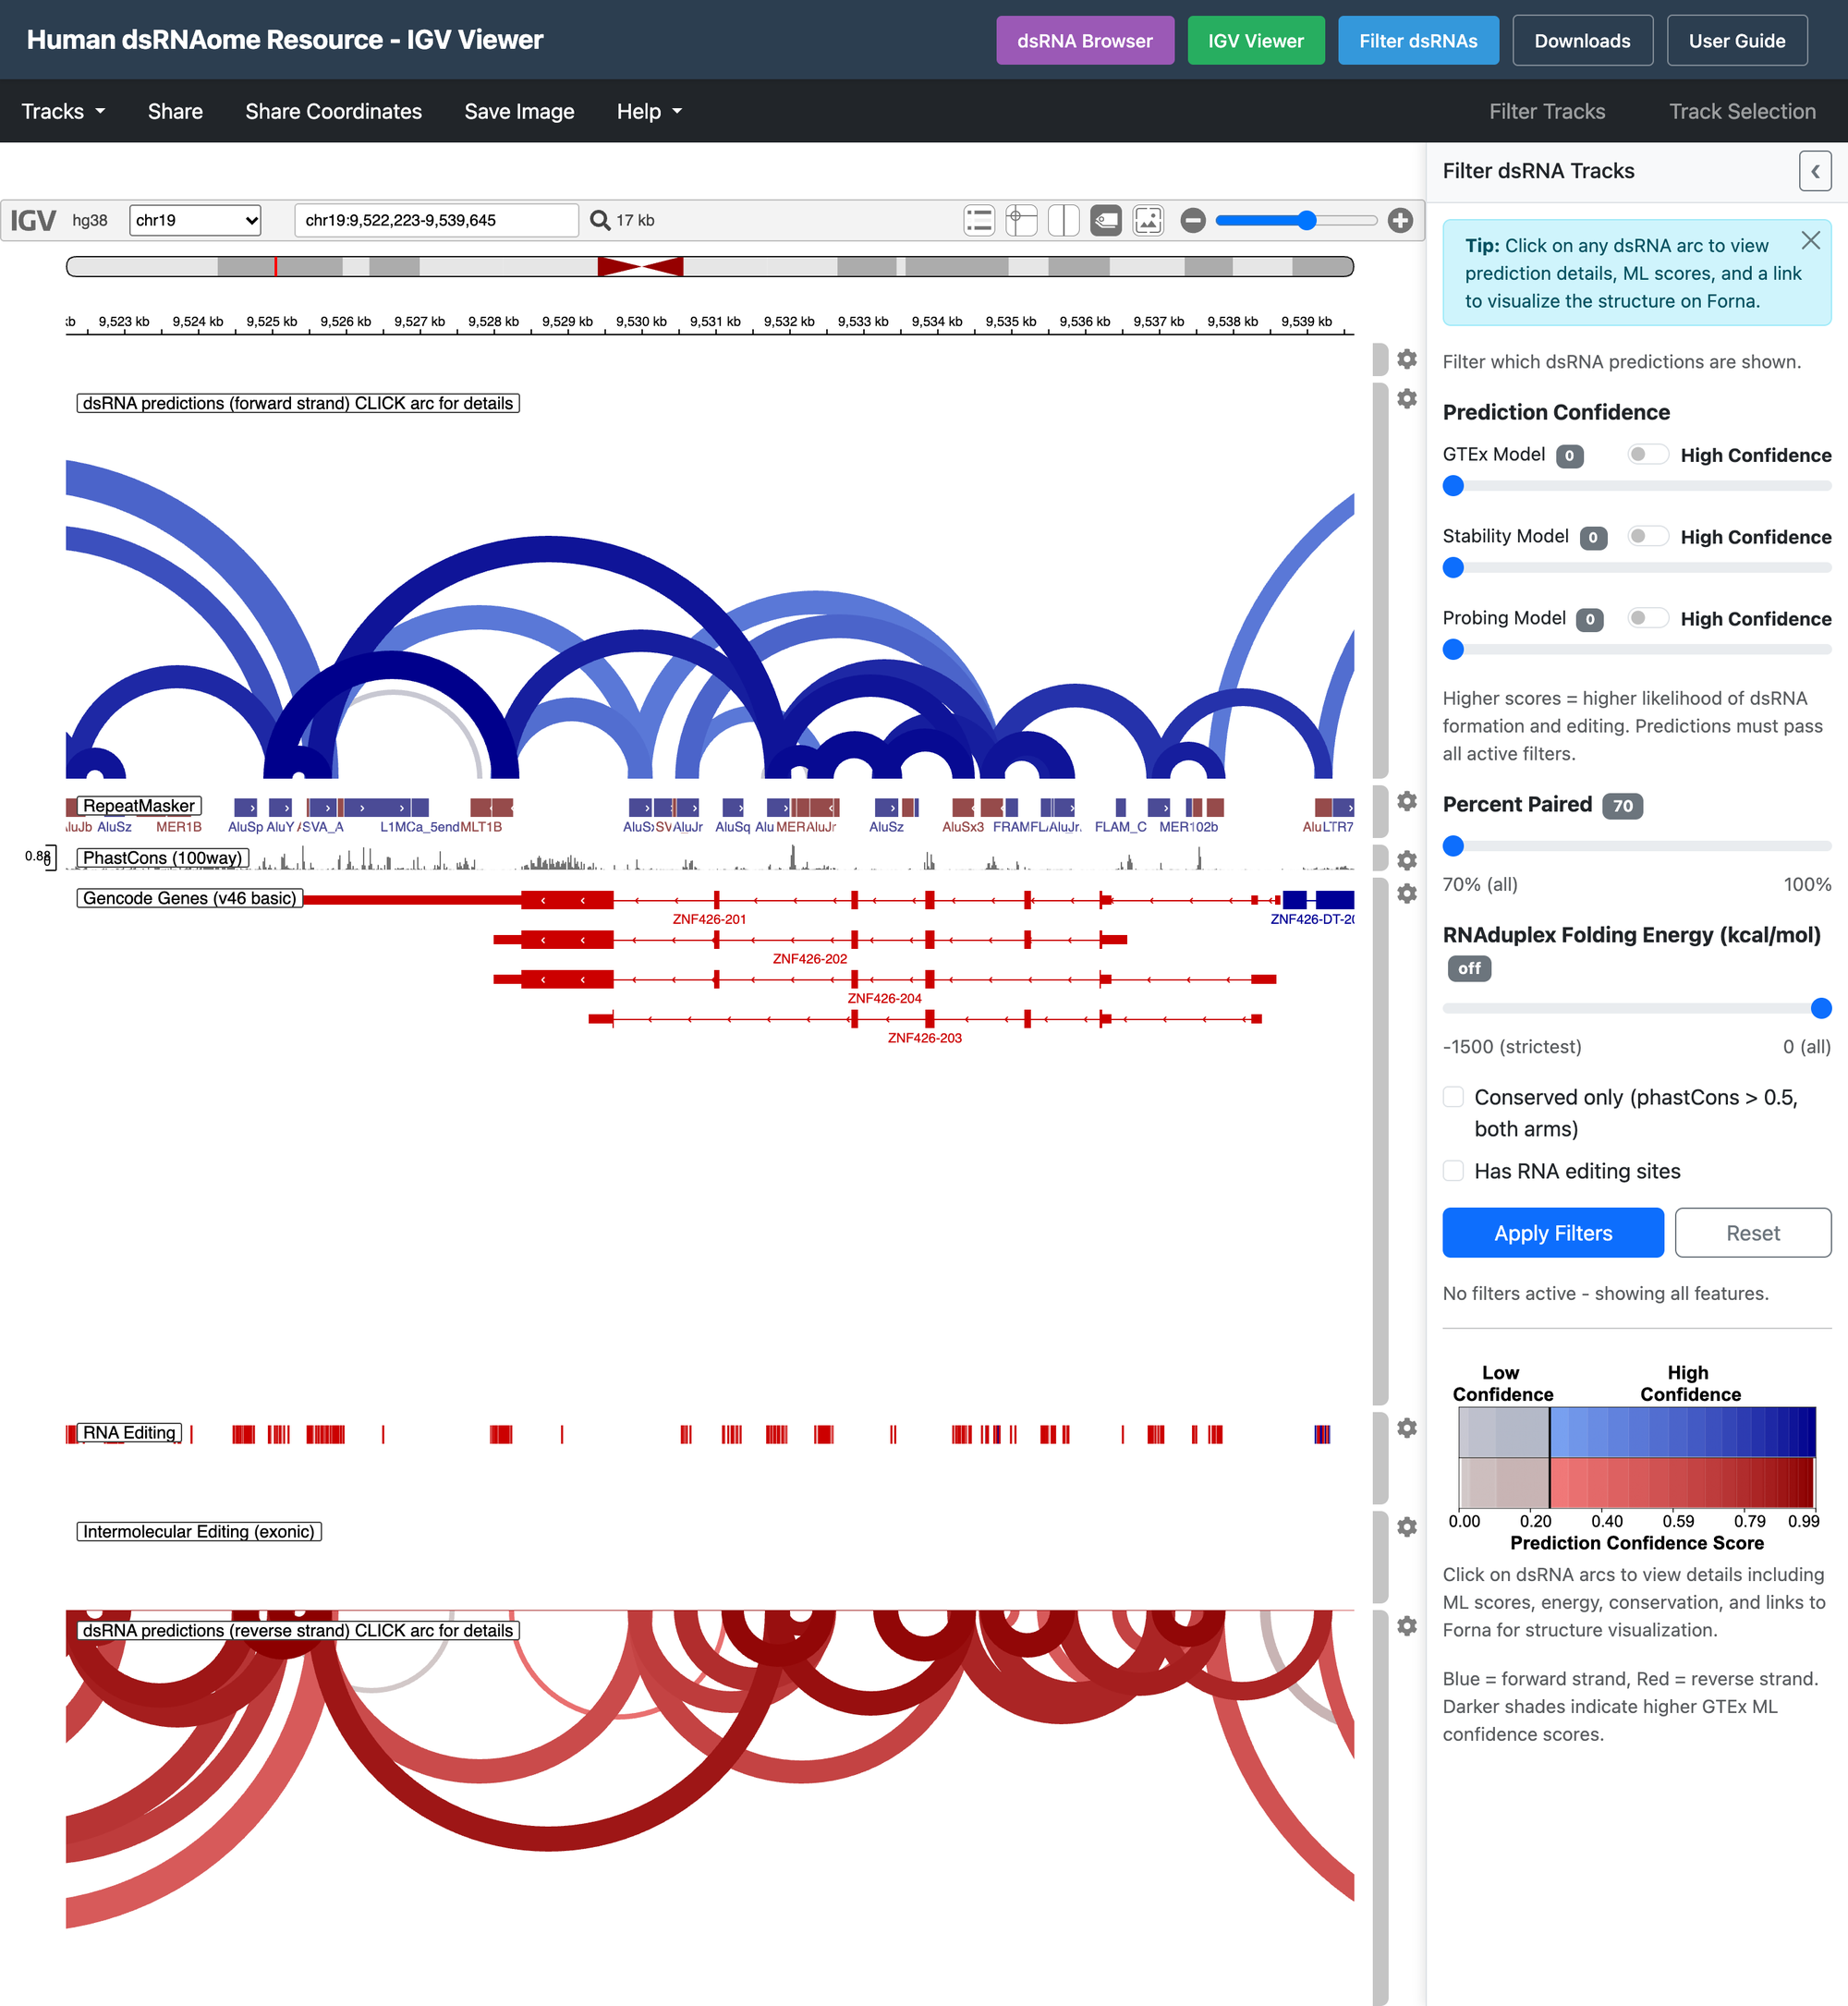
ZNF426in the locus box. Press Enter. Also accepted: an Ensembl ID (ENSG00000130270), coordinates (chr19:9,523,223-9,538,645), or two loci separated by a space.You should see: the gene ±1 kb with about 60 arcs. Only high-confidence predictions are drawn by default; untick High confidence only in the sidebar to see all 63. -
Read the arcs. One arc = one dsRNA, the arcs show which regions (arms) are predicted to form a dsRNA. Blue, opening up = forward strand. Red, opening down = reverse strand. Darker = higher GTEx score (key at the bottom of the sidebar).
 1
2
3
4
5
6
7
8
- Locus box: gene, Ensembl ID or coordinates
- Zoom slider (chromosome menu is left of the locus box)
- Ideogram: red box = where you are; click to jump
- Ruler: drag across a region to zoom into it
- Forward-strand dsRNAs: blue arcs opening up; darker = higher GTEx score
- Context tracks: RepeatMasker, PhastCons, GENCODE genes, RNA editing
- Reverse-strand dsRNAs: red arcs opening down
- Filter sidebar
- Move. Drag a track to pan; drag on the ruler, or use + / −, to zoom. Dense tracks stop drawing when zoomed out too far (limits).
-
Click an arc. The popup lists its scores, energy, % paired, genes, edits per arm, phastCons, and links to the dsRNA Browser and FORNA.
 1
2
- Click the arc (here dsRNA 2551194, isolated with the Percent Paired filter at 96%)
- Its popup: ID, links to the dsRNA Browser and FORNA, energy, % paired, loop length, the three model scores, genes, edits, phastCons
You should see: the dark red arc at chr19:9,524,968–9,525,786 isdsRNA_chr19_2551194: 96% paired, −590 kcal/mol, GTEx 0.97. -
Arrange the tracks for your gene. ZNF426 is on the minus strand, so its dsRNAs are the red arcs at the bottom. Menu bar → Swap strands ⇅ puts them on top, opening upward, like a plus-strand gene. To drop the forward-strand track altogether: Track Selection → untick dsRNA predictions (forward strand) → Apply. The same panel switches on the tracks that are off by default (PhastCons 20-/17-way, all intermolecular candidates, eCLIP).
 1
2
- Untick dsRNA predictions (forward strand)
- Apply
More ways to arrange tracks
Hover a track name for its gear menu: height, colour, expanded or collapsed (useful for the gene track), remove. Drag the handle at a track's left edge to reorder. Tracks → URL... / Local File... loads your own data in any igv.js format (BED, bigWig, GFF, BAM); the file stays in your browser. Share links record which tracks are on (tracks=). -
Change which predictions are drawn. The default is High confidence only: the 2,458,476 predictions that pass at least one model's cutoff. Untick it and Apply Filters for all 5.1 million. To tighten instead, raise a slider (GTEx High Confidence snaps it to 0.25); Sliders combine with sets AND or OR when more than one is up. Reset returns to the default.
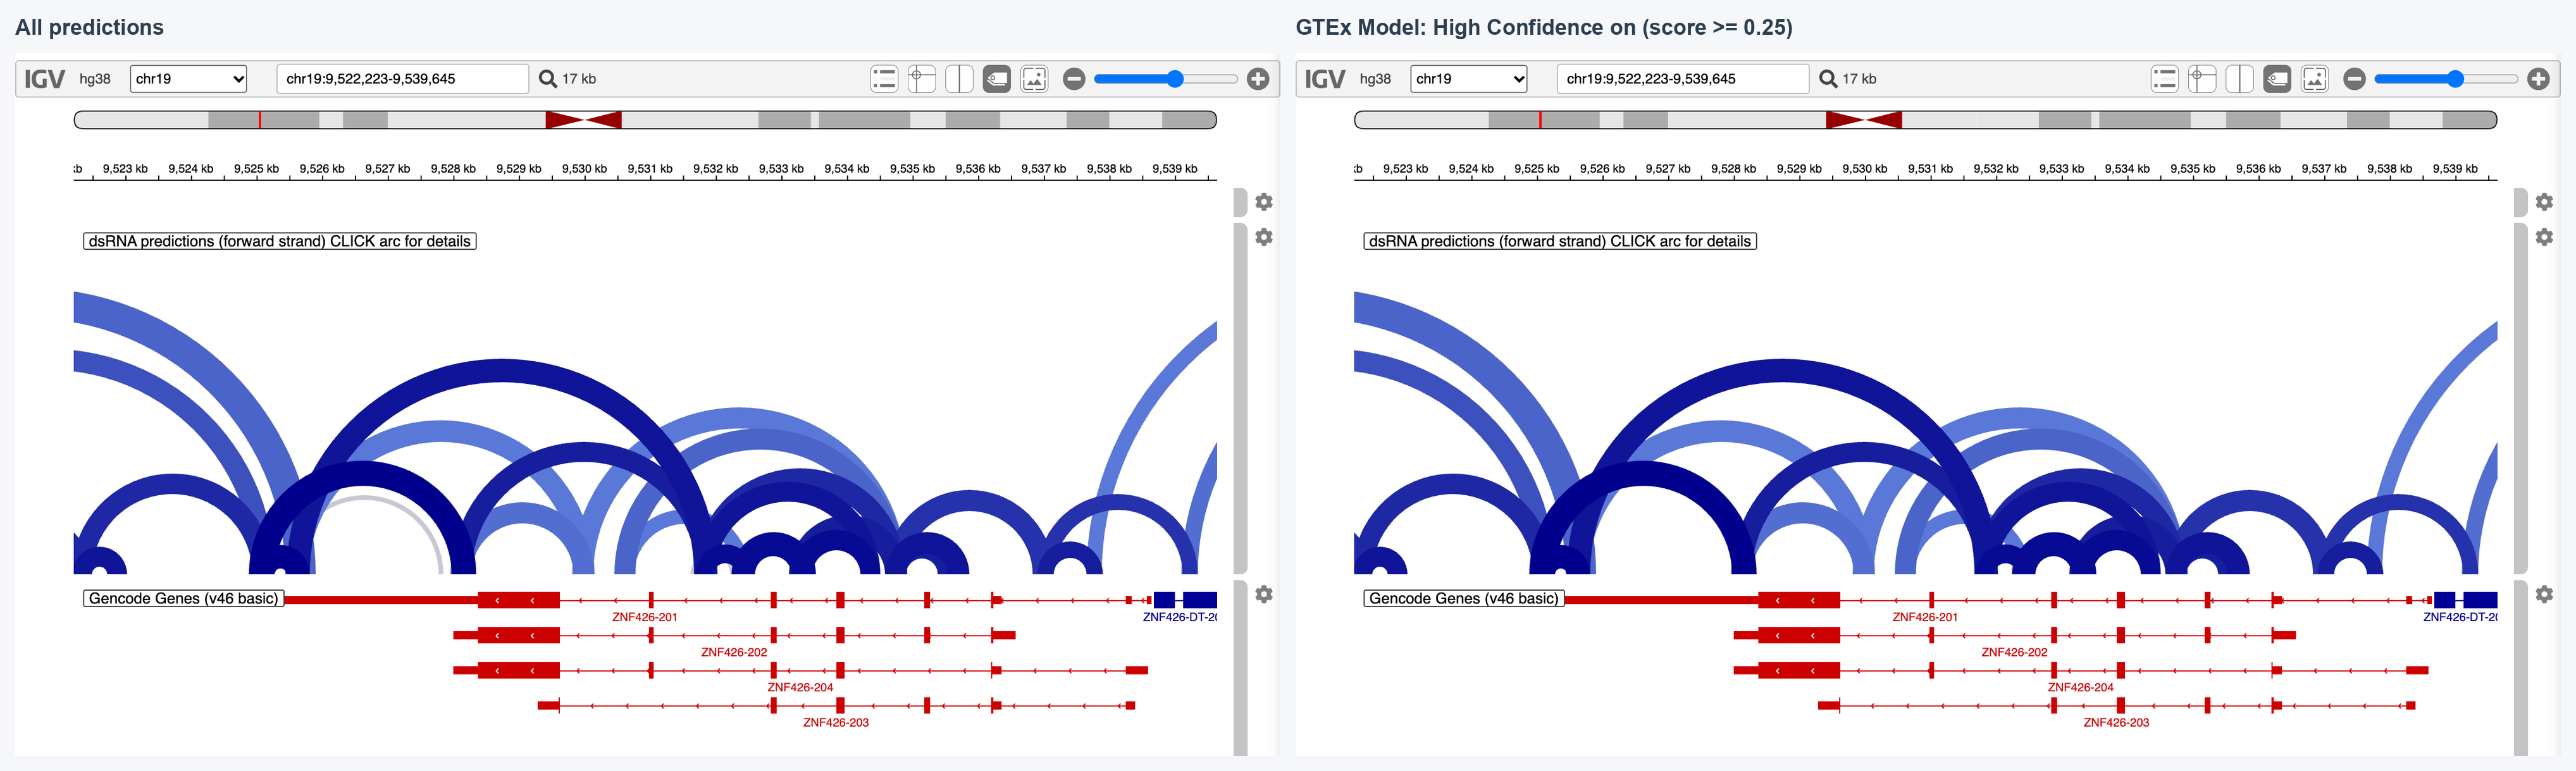
The reverse-strand track from step 5. Left: all predictions (High confidence only unticked). Right: GTEx High Confidence. The pale, low-scoring arcs are gone; the dark ones stay.
Next: evaluate the arc you clicked in the dsRNA Browser, or find every dsRNA like it genome-wide.
2. Find long, highly paired dsRNAs
Filter dsRNAsKeep only dsRNAs long and paired enough for sensors like MDA5 or PKR, then export the list.
-
Gene name(s) box: type
ZNF426. Several genes:MYC, ADAR, IFIH1. Empty box = whole genome. -
Click Apply Filters, then open the Summary tab.
You should see: Matched N dsRNAs (x% of 5,134,754 total in DB), the filters applied, and counts by editing and repeat status.
 1
2
3
4
- Gene name(s) or Ensembl ID(s)
- Apply Filters
- Summary tab: matches, filters applied, editing and repeat breakdown
- Download...: every matching row as CSV, Parquet or BEDPE
-
Set Percent Paired to 96% and Length to 300–5000 bp. Apply Filters again.
You should see (ZNF426): one dsRNA, chr19:9,524,968–9,525,786 (−), 96% paired, −590.7 kcal/mol.

Why 96% and 300 bp?
MDA5 filaments need long, continuous duplex; 96% pairing on arms ≥ 300 nt keeps structures with few bulges. The ZNF426 duplex is ~290 bp with six A-C mismatches, one A-G, a two-pair internal loop and one bulge (paper Figure S5). Lower pairing to 90% to admit Alu pairs, the bulk of the dsRNAome. Add Conserved, >1 editing site per arm or the High Conf boxes to tighten further. - Open the Data Table tab. Per row: View on IGV, View Structure (FORNA). All columns adds per-arm coordinates, conservation, sequence and structure.
- Download... → format → columns → Download. CSV, CSV.gz, Parquet or BEDPE. The file has every matching row, not just the rows on screen.
Gotchas (3)
- Apply before Download. The download reads the sidebar as it is now, not as it was at the last Apply.
- Table capped at 1,000 rows (raise under "Maximum results in table", max 50,000). The search box above the table only searches those rows.
- Locus box bounds the i-arm only. For "both arms inside", download and filter on the per-arm coordinates.
Next: click View on IGV on the row, then evaluate it in the dsRNA Browser.
3. Evaluate one candidate
dsRNA BrowserDo probing and editing data support this dsRNA, and does the duplex explain the data better than each arm folding alone?
MAVS 3189903: 18 probing experiments, best ROC-AUC 0.87, 44 editing sites, GTEx 0.93. ZNF426 2551194: GTEx 0.97 and 60 editing sites, but one in-vitro experiment at ROC 0.62.
-
Open the dsRNA. From an IGV popup (View in dsRNA Browser), the dsRNA ID box (exact number), Gene Search (partial match), or a link:
https://dsrna.chpc.utah.edu/api/dsrna-browser/#dsRNA_3189903. First visit: wait for the loading bar in the header to finish.Reading the table
Sorted by Best Reactivity (powered): the ROC-AUC of the best validation that passes the paper's power filter (detection power ≥ 0.5), where "best" means the highest lower 95% confidence bound, so a 1.000 from a few nucleotides ranks below a 0.87 from fifty. Hover the value for its CI and nucleotide counts. Rows with no powered validation show their unfiltered ROC in grey brackets with ⚡ and sort last. Green/red cells: ML GTEx and Stability above/below cutoff; ML Probing red → yellow → green; reactivity green ≥ 0.7, orange 0.5–0.7, red below. Power-filtered only keeps the ~15% of rows with a powered validation.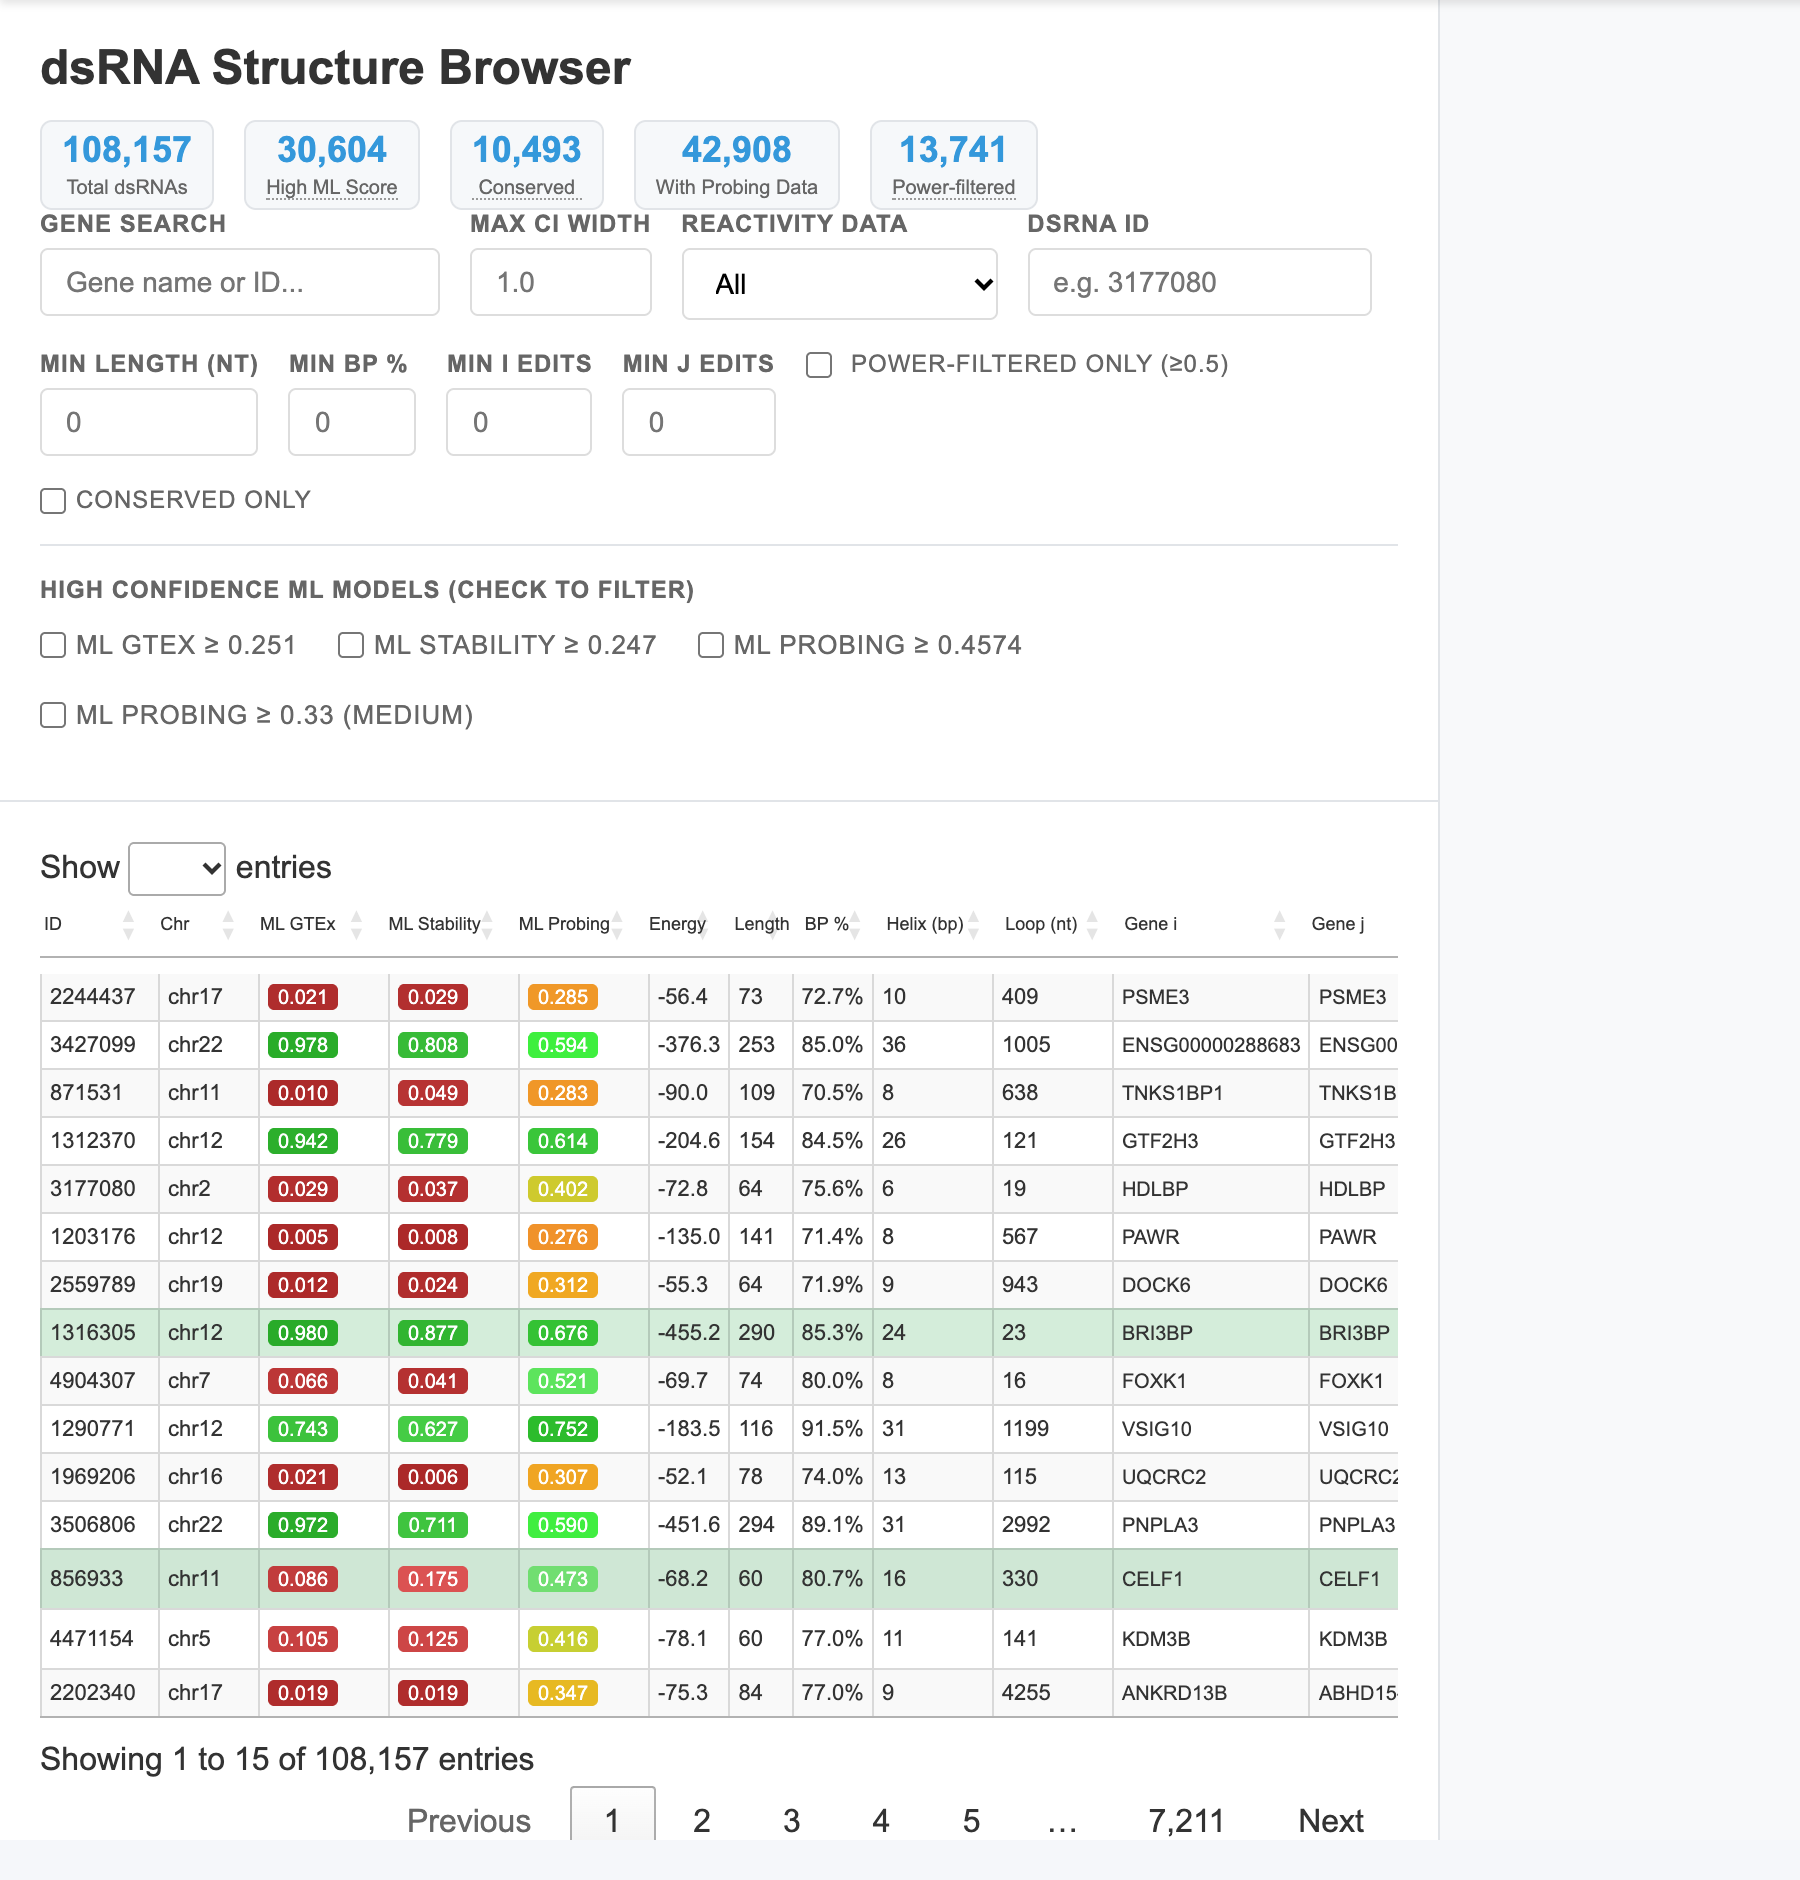 1
2
3
4
- Counts: total, high ML score, conserved, with probing data, power-filtered
- Gene Search (partial match, either arm)
- dsRNA ID (exact)
- Power-filtered only
-
Pick the experiment. Select Reactivity Experiment lists each as
name [i:ROC, j:ROC]; the best is ★ and preselected. -
Read the structure. Fill = reactivity, pale (paired) to red (unpaired); grey = no data. Green outline = editing site, rare to common. Hover a base for its values.
A supported duplex looks like: pale stems, red loops. Red inside a long stem means the data disagree there.
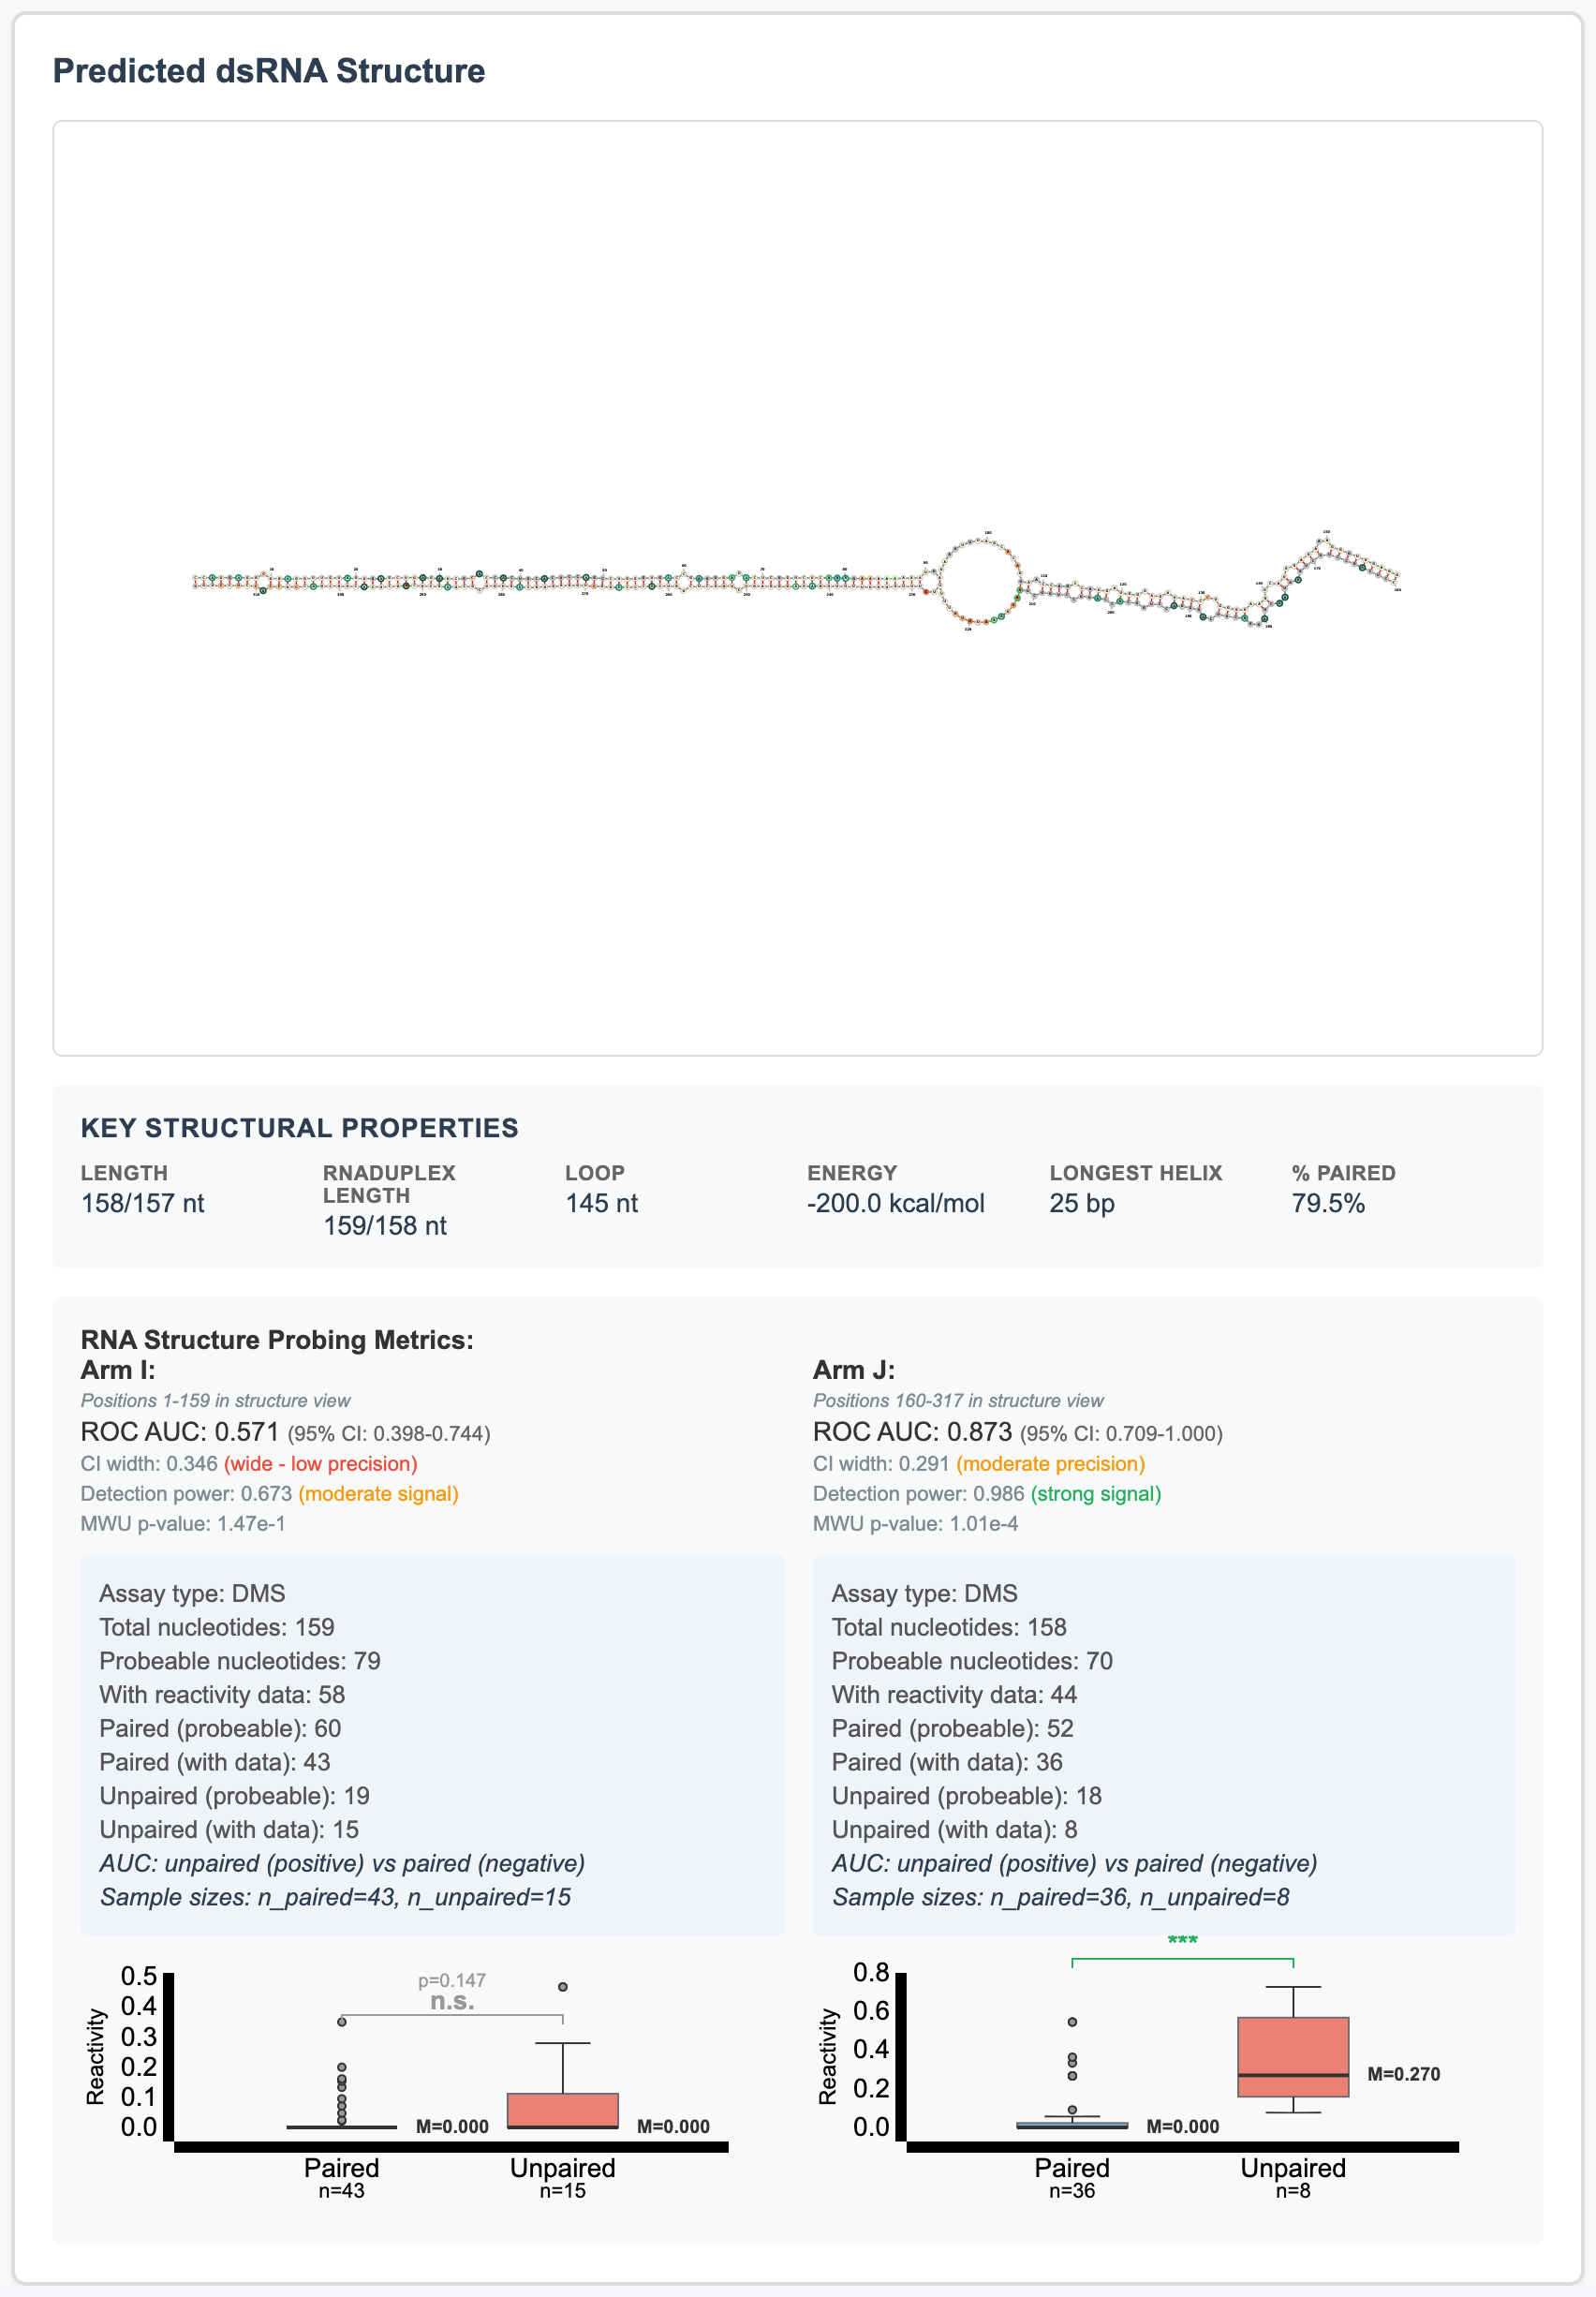
The structure panel for dsRNA 3189903: the drawing (drag to pan, scroll to zoom), Key Structural Properties, and the per-arm metrics with their paired-vs-unpaired box plots. - Check the numbers. Per arm under the structure: ROC AUC (0.5 = chance, > 0.7 = good), 95% CI (width < 0.1 = precise), power, p-value. ⚠️ = few data points or a perfect 1.000; ⚡ = low power. Then RNA Structure Probing Summary: ★ marks dsRNA vs RNAfold winner per arm (difference ≥ 0.05). And RNA Editing Sites: sites on both arms = strongest evidence the arms pair in cells.
- Keep it. Download dsRNA Summary (ZIP: FASTA, DBN, CT, BED, probing CSV, SVGs, summary page) or Print Summary (PDF-ready).
Next: Open in IGV Browser → under Genomic Context, or download data.
4. Download data
Filter dsRNAs Downloads dsRNA Browser| You want | Where | You get |
|---|---|---|
| A filtered subset | Filter dsRNAs → Download... | CSV, CSV.gz, Parquet, or BEDPE (38 fixed columns, no header, 0-based). Every matching row. dsRNAscan_YYYY-MM-DD.<ext> |
| The whole dsRNAome | Downloads → Download the main dataset | Parquet and BEDPE of all 5,134,754; high-confidence subsets (2,458,476 any model; 1,509,138 all three); column dictionary. |
| Tracks for IGV desktop | Downloads → Browser tracks & pipeline files | .bp arc files, GFF3 annotations, the files the online viewer uses. |
| One dsRNA's structure files | dsRNA Browser → Download dsRNA Summary | ZIP: FASTA, DBN, CT, BED, probing CSV, SVGs, JSON, summary HTML. |
| Code | Downloads → Recipes | pandas / polars / DuckDB loaders; bedtools overlaps with m6A, splice sites, eCLIP. |
Pick: Parquet for code (smallest, typed), CSV for a spreadsheet, BEDPE for bedtools.
Reference: IGV Viewer
The IGV Viewer is igv.js with the dsRNAome tracks preloaded and a filter sidebar added. Standard igv.js behaviour (track menus from the gear icon, drag to reorder tracks, right-click options) all works.
Tracks and zoom limits
Every feature track stops drawing beyond a maximum window so that dense data do not stall the page. When a track is blank, check the zoom first.
| Track | Default | Shows | Source | Drawn up to |
|---|---|---|---|---|
| dsRNA predictions (forward strand) | on | Arcs, blue, opening upward; shade = GTEx score | dsRNAscan, this resource | 1 Mb |
| dsRNA predictions (reverse strand) | on | Arcs, red, opening downward; shade = GTEx score | dsRNAscan, this resource | 1 Mb |
| RepeatMasker | on | Repeats (Alu, LINE, LTR, simple), collapsed | UCSC rmsk, hg38 | 1 Mb |
| PhastCons (100way) | on | Conservation across 100 vertebrates, 0–1 | UCSC | 500 kb |
| PhastCons (20way) | off | Conservation across 20 mammals | Broad igvdata | 500 kb |
| PhastCons (17way) | off | Conservation across 17 primates | UCSC | 500 kb |
| Gencode Genes (v46 basic) | on | Transcripts, expanded, labelled by transcript name (e.g. SON-202) | GENCODE v46 basic set | 2 Mb |
| RNA Editing | on | One tick per A-to-I site, collapsed | REDIportal v3, 15.7 M sites | 150 kb |
| Intermolecular Editing (exonic) | on | Editing clusters explained by pairing between two different transcripts (exonic subset) | this resource | any |
| Intermolecular Editing Candidates (all) | off | All strand-specific intermolecular candidates | this resource | any |
| eCLIP binding sites | off | RBP binding peaks, one row per protein / cell line / replicate | ENCODE eCLIP | 75 kb |
Filter sidebar
The sidebar hides arcs from both dsRNA tracks. Nothing changes until you click Apply Filters; Reset returns every control to its default and re-applies. Every active condition must pass, except that the three model sliders can be combined with OR.
| Control | Range | Keeps a dsRNA when |
|---|---|---|
| High confidence only (any model) | checkbox, on by default | GTEx ≥ 0.25 or Stability ≥ 0.25 or Probing ≥ 0.46: the paper's 2,458,476 high-confidence predictions. Untick for all 5,134,754. |
| GTEx Model + High Confidence | 0–1 (toggle = 0.25) | GTEx model score ≥ slider |
| Stability Model + High Confidence | 0–1 (toggle = 0.25) | Stability model score ≥ slider |
| Probing Model + High Confidence | 0–1 (toggle = 0.46) | Probing model score ≥ slider |
| Sliders combine with | AND (default) / OR | with more than one slider raised: pass every one (AND) or any one (OR) |
| Percent Paired | 70–100% (70 = off) | percent paired ≥ slider |
| RNAduplex Folding Energy | −1500–0 kcal/mol (0 = off) | energy ≤ slider (more negative = stricter) |
| Conserved only | checkbox | phastCons 100-way and 17-way > 0.5 on both arms (stricter than the table's filter, which uses the 100-way score only) |
| Has RNA editing sites | checkbox | at least one same-strand REDIportal site on either arm (looser than the paper's "edited", which needs more than one site on each arm) |
The sidebar can be hidden with the ❮ button or Filter Tracks in the menu bar; the IGV panel widens to fill the space.
Menu bar
- Tracks
- Local File... loads a track from your disk; URL... loads one from a web address. Indexed formats (.bed.gz + .tbi, bigWig, bigBed, BAM + .bai) stream only the region in view.
- Share
- Link to the current view (locus + tracks + filters), embeddable iframe, QR code.
- Share Coordinates
- A plain locus link in a prompt box for quick copying.
- Save Image
- SVG (vector) or PNG of the whole panel.
- Help
- The igv.js user guide, this guide, and the About box with version numbers.
- Swap strands ⇅
- Mirror the arc layout: reverse-strand arcs on top opening upward, forward-strand arcs at the bottom opening downward. For genes on the minus strand. Recorded in share links as
flip=1. - Filter Tracks
- Show or hide the filter sidebar.
- Track Selection
- Checklist of all built-in tracks with Select All / Unselect All / Apply.
Reference: Filter dsRNAs (table)
A Shiny app over a SQLite database of all 5,134,754 predictions. Filters are combined with AND; within the gene box, the tokens are ORed.
Search
- Gene name(s) or Ensembl ID(s)
- Symbols match either arm's gene exactly (case-insensitive).
ENSGIDs (version suffix allowed) match the i-arm's gene ID. Separate several with commas, semicolons, spaces or new lines. - Locus
chr4,chr4:1,000,000orchr4:1,000,000-2,000,000. Keeps dsRNAs whose i-arm starts at or after the start and ends at or before the end. Unparseable text is ignored without a message, so check the Summary tab's "Filters applied" list.- Strand
- Any, +, or −.
Filters
| Section | Control | Keeps a dsRNA when |
|---|---|---|
| Conservation | Conserved (phastCons > 0.5, both arms) | phastCons 100-way > 0.5 on both arms |
| RNA Editing | >1 editing site per arm, either strand | stranded + unstranded sites > 1 on each arm |
| Same Strand (i / j), Either strand (i / j) | the named count ≥ the number typed (0 = off) | |
| Repetitive Elements | All / Repetitive only / Non-Repetitive only | an arm overlaps (or no arm overlaps) a RepeatMasker element |
| Prediction Confidence | GTEx, Stability, Probing sliders; High Conf boxes | score ≥ slider. High Conf moves the slider to 0.25 / 0.25 / 0.46 |
| Percent Paired | 70–100% | percent paired ≥ value (70 = off) |
| Length (i or j) | 25–5000 bp range | at least one arm's length is inside the range |
| RNAduplex Folding Energy | −1500–0 | energy ≤ value (0 = off) |
| Maximum results in table | 100–50,000 (default 1,000) | rows rendered in the table; does not affect downloads or the Summary counts |
Tabs
- Data Table
- Essentials or All columns; client-side search, sort and per-column filters over the loaded rows; "View on IGV" and "View Structure" links per row.
- Summary
- Match count and percentage of the database, the filters applied, and counts by editing status, repetitive status, and the two crossed.
- Density Plots
- Arm length, longest helix, percent paired, loop length and folding energy, split by editing status and repetitive status. Drawn from a random sample of up to 20,000 matches, so distributions are representative even when the table is capped.
- Box Plots
- The same five variables as box plots by editing status, with n in the labels.
Download dialog
Format: CSV, compressed CSV, Parquet, BEDPE (default), compressed BEDPE. Columns: one checkbox per database column, preselected to the Essentials set; quick links Essentials / All / None. BEDPE ignores the column picker: columns 1–10 are standard BEDPE (chr1, start1, end1, chr2, start2, end2, name, score, strand1, strand2) and 11–38 carry every attribute, with no header row. Large downloads can take 30 seconds or more; a progress bar shows the stage.
Reference: dsRNA Browser
A single-page app that loads its database into the browser once (cached for the session), then shows a filterable table and a detail view for the selected dsRNA. The database is the ~108,000 dsRNAs whose arms overlap exons; 13,789 of them have probing data that passes the paper's power filter (≥ 0.5), and those are the ones whose ROC-AUC values are worth reading closely.
Table columns
Click any header to sort. The sort and page size persist between visits. The default order is Best Reactivity (powered): among experiment-arms with detection power ≥ 0.5 (the paper's power filter), the validation with the highest lower 95% confidence bound on its ROC-AUC. Ranking by the bound rather than the ROC itself keeps a perfect 1.000 from a handful of nucleotides from outranking a well-measured 0.87. A green left border marks rows where one experiment has both arms powered with ROC > 0.7; a red border marks both arms powered with ROC < 0.5 (well-measured, and the duplex is contradicted).
Filters
- Gene Search
- Substring match on the gene name or Ensembl ID of either arm.
- dsRNA ID
- Exact numeric ID.
- Max CI Width
- Keep rows whose best-ROC confidence interval is at most this wide (rows without a CI are kept).
- Reactivity Data
- All / Both Arms / Any Arm / No Reactivity Data.
- Min Length, Min BP %, Min i Edits, Min j Edits
- Numeric minimums.
- Power-filtered only (≥0.5)
- At least one experiment-arm with detection power ≥ 0.5; about 15% of the table.
- Conserved only
- All phastCons scores > 0.5.
- High Confidence ML Models
- GTEx ≥ 0.251, Stability ≥ 0.247, Probing ≥ 0.4574, or Probing ≥ 0.33 (medium, includes GRIA2-like structures). Ticked boxes are ANDed.
When any filter is active a red banner shows "showing X of Y entries" with a Clear All Filters button.
Detail view, top to bottom
- Select Reactivity Experiment: dropdown; best experiment starred and preselected.
- Reactivity legend: low → high colour bar, grey = no data, green outline = editing site.
- Predicted dsRNA Structure with Key Structural Properties (arm lengths, RNAduplex-trimmed lengths, loop, energy, longest helix, % paired) and the per-arm RNA Structure Probing Metrics.
- Genomic Context: location, "Open in IGV Browser", and an embedded IGV view of the dsRNA ±100 nt with its base pairs and GENCODE genes.
- RNA Editing Sites (when present): table of REDIportal sites with sample and tissue counts; link to REDIportal.
- Local Folding (RNAfold): each arm folded alone, with MFE energy and, when an experiment is selected, the same probing metrics.
- RNA Structure Probing Summary: dsRNA structure vs RNAfold ROC per arm; ★ marks the better fit.
Interpreting scores
A prediction in this resource means dsRNAscan found an inverted repeat that can form a duplex of at least 25 bp at ≥ 70% pairing. Everything else is a layer of evidence on top of that. The layers are independent, so a candidate supported by two or three of them is far more likely to be a real, cellular dsRNA than one supported by none.
The three machine-learning scores
| Score | Trained to predict | Uses | High-confidence cutoff | Performance |
|---|---|---|---|---|
| GTEx Model | Whether the dsRNA is edited by ADAR (both arms) | Structure features plus GTEx tissue expression | ≥ 0.25 (0.251) | AUC 0.961; 93.8% sensitivity at the cutoff |
| Stability Model | Whether the dsRNA is edited by ADAR | Structure and thermodynamic features only, no expression | ≥ 0.25 (0.247) | AUC 0.914; 96.4% sensitivity |
| Probing Model | Whether chemical probing data agree with the predicted structure | Structure features; score min-max normalised to 0–1 | ≥ 0.46 (0.4574); 0.33 = medium | AUC 0.847 |
The IGV Viewer and Filter dsRNAs round the cutoffs to two decimals; the dsRNA Browser shows the exact values. They select the same dsRNAs. 2,458,476 predictions pass at least one model's cutoff (the paper's high-confidence set) and 1,509,138 pass all three. The GTEx and Stability models agree on 1.7 million; the Stability model adds about 471,000 that the GTEx model misses (mostly dsRNAs with little GTEx expression) and the GTEx model adds about 58,000 (Figure 3C of the paper). Both subsets are on the Downloads page as BEDPE files.
What a high score does not mean. The GTEx and Stability models were trained on ADAR editing, so a high score says "this looks like the dsRNAs ADAR edits", not "this is immunogenic" or "this is functional". A dsRNA with no expression in GTEx tissues can score low on the GTEx model and high on the Stability model; that is a difference in expression, not in structure. Use the Stability model for tissues or conditions GTEx does not cover.
Experimental layers
- Editing sites
- REDIportal v3 A-to-I sites (9,642 GTEx samples, 54 tissues; 9,681 TCGA samples). Sites on both arms are the strongest signal that the two arms pair in vivo. Sites on one arm only may come from a different partner. Sites on the opposite strand still count (the "unstranded" columns): the same inverted repeat forms the same duplex on either strand, so editing of one strand is taken as evidence for the mirror-image prediction on the other, which is unedited only because that strand is not expressed. See the glossary.
- Probing ROC-AUC
- From DMS, SHAPE and related experiments: how well reactivity separates predicted-unpaired from predicted-paired nucleotides. Before any filtering, most of the 189,000 arm-level AUCs sit near 0.5; applying the paper's power filter (≥ 0.5) keeps 35,000 of them and the distribution splits into supported (> 0.6) and contradicted (< 0.4) structures. Above 0.7 with a CI width under 0.1 on both arms is strong agreement. Always read the n: a ROC of 1.0 from six nucleotides means little; a power-filtered validation has a median of 56 paired and 17 unpaired nucleotides with data.
- Conservation
- phastCons 100-way (vertebrates) and 17-way (primates), per arm. Most of the dsRNAome is Alu-derived and primate-specific, so a conserved dsRNA (both arms > 0.5) is unusual and worth a closer look.
- Repeat status
- Over 90% of predictions involve a repeat, mostly inverted Alu pairs. Non-repetitive dsRNAs are rarer and more often conserved.
Rules of thumb
- Immune-sensor candidates: long arms (≥ 300 nt), ≥ 90–96% paired, strongly negative energy. Then check the Probing Model and, in the dsRNA Browser, whether any experiment supports the duplex.
- ADAR substrates: GTEx or Stability high confidence plus editing on both arms.
- Regulatory candidates in a specific gene: look at where the arms fall (3′ UTR, intron, exon) in the IGV gene track, and whether an eCLIP peak overlaps an arm.
- Prioritising a long list: sort by the number of models calling high confidence (the "# Models High-Conf" column under All columns), then by editing counts.
Glossary
- dsRNAscan
- The pipeline that produced the predictions: overlapping 10 kb windows scanned for inverted repeats that RNAduplex folds into a duplex of ≥ 25 bp at ≥ 70% pairing. Code at github.com/Bass-Lab/dsRNAscan.
- i-arm / j-arm
- The two halves of the inverted repeat. i is upstream (lower genomic coordinate); j is downstream. On the minus strand the j-arm is transcribed first.
- Loop
- The genomic distance between the arms. It can be a few nucleotides or many kilobases; long loops are typical of intronic Alu pairs.
- Percent paired
- Fraction of nucleotides in the predicted duplex that are base-paired (including G-U).
- Longest helix
- The longest run of consecutive base pairs with no bulge or mismatch.
- Energy
- RNAduplex free energy of the duplex in kcal/mol. More negative = more stable. Scales with length, so compare energies between dsRNAs of similar size.
- RNAduplex length
- Arm lengths after RNAduplex trims unpaired ends; shown in the dsRNA Browser next to the original lengths.
- Edited (paper definition)
- A dsRNA is called edited when more than one REDIportal site falls on each arm, counting sites on either strand. 1,236,563 of the 5.1 million predictions are edited by this definition, over 99% of them in repetitive elements.
- Stranded vs unstranded editing sites
- Stranded sites are REDIportal sites on the same strand as the dsRNA; unstranded sites are on the opposite strand. Both count as evidence, for this reason: an inverted repeat is complementary on both strands, so every prediction on the plus strand has a mirror-image prediction on the minus strand. If one strand is transcribed and clearly edited, the opposite-strand prediction would fold the same way and be edited too, were that strand expressed. Its lack of same-strand sites reflects expression, not structure, so the resource credits it with the editing seen on the other strand.
- Intermolecular editing
- Editing clusters that cannot be explained by a duplex within one transcript and are explained by pairing between two different transcripts.
- ROC-AUC
- Area under the receiver operating characteristic curve. Here: the probability that a randomly chosen predicted-unpaired nucleotide has higher reactivity than a randomly chosen predicted-paired one.
- CI width
- Width of the 95% confidence interval on the ROC-AUC (Hanley & McNeil 1982). Narrow = many data points.
- Detection power
- Estimated power to detect the observed paired/unpaired reactivity difference given the sample size.
- phastCons
- Per-nucleotide probability of being in a conserved element, 0–1, averaged over an arm.
- FORNA
- The ViennaRNA structure-drawing tool used in the dsRNA Browser and linked from IGV and the table.
- BEDPE
- Paired-end BED: one line per dsRNA with both arms' coordinates (0-based, half-open), the format bedtools pairtobed reads.
FAQ and troubleshooting
Arc or editing track is empty
Cause: zoomed out past the track limit (arcs 1 Mb, editing 150 kb, eCLIP 75 kb), or a filter hides everything.
Fix: type a gene name or drag a smaller region on the ruler. Check the status line under Apply Filters; click Reset.
"Unrecognized locus" for my gene
Cause: names come from GENCODE v46 symbols and ENSG IDs only; aliases and ENST IDs are not searched.
Fix: use the current HGNC symbol (IFIH1, not MDA5), an ENSG ID, or coordinates.
The table found my gene but not the dsRNA I saw in IGV
Cause: the 1,000-row cap, a leftover filter, or the gene is on neither arm.
Fix: raise "Maximum results in table"; check the Summary tab; search the Locus box with the coordinates from the IGV popup.
dsRNA Browser can't find an ID that IGV links to
Cause: the browser holds ~108,000 exon-overlapping dsRNAs, not all 5.1 million.
Fix: none needed; its scores and coordinates are in IGV and the table.
"Structure Too Large for FORNA Visualization"
Cause: an arm is over 1,000 nt.
Fix: metrics, editing table and downloads still work. Open the .dbn or .ct from Download dsRNA Summary in RNAstructure or VARNA.
ROC-AUC is 1.000 but flagged
Cause: very few probeable nucleotides with data.
Fix: read "With reactivity data" and the CI width; pick another experiment from the dropdown, or treat as inconclusive.
Download doesn't match the table
Cause: the download reads the sidebar now; the table shows the last Apply.
Fix: click Apply Filters, confirm the Summary tab, then download.
Low GTEx score but high Stability score
Cause: the GTEx model includes tissue expression; a gene with little GTEx expression scores low on it regardless of structure.
Fix: use the Stability model for cell lines, tumours or tissues GTEx does not sample.
Can I load my own data next to the predictions?
Yes. IGV Viewer → Tracks → URL... or Local File..., any igv.js format. Large files: use an indexed format (bgzipped BED/GFF + .tbi, bigWig, bigBed, BAM). Nothing is uploaded to the server.
IGV is slow when zooming or dragging
Cause: every loaded track redraws on each step; the expanded gene track and eCLIP are heaviest.
Fix: hide unused tracks in Track Selection; collapse the gene track from its gear menu.
Citing and contact
If you use the dsRNAome in published work, please cite:
Andrews RJ and Bass BL. dsRNAscan maps human dsRNAome revealing conservation, intermolecular dsRNA, and determinants of ADAR dependency.
Questions, bug reports and feature requests: ryan.andrews@biochem.utah.edu, bbass@biochem.utah.edu.
The original user guide published with the paper (Document S1) is available as a PDF. It describes an earlier version of the IGV page; this page is the current guide.